





A clean-label claim is only as credible as the documentation behind each ingredient. A supplier’s brochure, natural-sounding product name, or statement that an ingredient is “free from artificial additives” does not establish how the material was made, what it contains, or whether it fits the intended market. Verification starts by treating the document set as one connected record: the ingredient specification, certificate of analysis, safety data, origin statement, allergen declaration, processing description, regulatory statement, and traceability records should describe the same material without unresolved contradictions.
The first review should establish a precise identity. Trade names are often broad enough to cover several grades, concentrations, carriers, or physical forms. “Natural vanilla flavor,” “citrus extract,” “fermented thickener,” or “plant-derived surfactant” can each refer to materially different compositions. The documentation should state the common or chemical identity where appropriate, botanical name for plant materials, plant part, extraction medium, concentration or standardization basis, and any carrier, preservative, processing aid, or anticaking agent present in the supplied form.
A document that identifies only the headline ingredient leaves a major gap. For example, a rosemary extract may be supplied in oil, ethanol, glycerin, maltodextrin, or another carrier. The extract itself may suit a clean-label formulation while the carrier changes the finished-product declaration, allergen review, flavor profile, solubility, or regional suitability. The same issue arises with powders that contain flow agents, liquid flavors diluted in solvents, peptide solutions stabilized with buffers, and natural pigments dispersed in emulsifiers.
Build the review around the material actually delivered
Start with the current specification and link it to the exact supplier code, grade, manufacturing site, and revision date. A generic technical data sheet is useful background, but it should not be accepted as the controlling document when a grade-specific specification exists. Revision control matters because a small document update can reflect a changed carrier, altered test method, wider microbiological limit, different source country, or revised shelf life.
The specification should provide measurable acceptance criteria rather than descriptive claims alone. Relevant parameters vary by ingredient class:
- For antioxidants and preservatives, review active-content range, solvent residues where relevant, moisture, oxidation indicators, microbial limits, and the method used to measure potency.
- For hydrocolloids such as gums, pectins, starches, and fibers, compare viscosity method, test concentration, temperature, shear conditions, pH, particle size, hydration behavior, and microbial specification. A viscosity value without method conditions cannot reliably predict behavior in a finished formula.
- For flavors, essential oils, and botanical extracts, examine identity markers, refractive index or density where suitable, chromatographic profile where available, declared solvents, flash point when applicable, and limits for contaminants associated with the source material.
- For cosmetic actives, assess assay, solvent system, pH, microbial controls, preservative system, residual monomers or reagents where applicable, and stability conditions. A high assay does not demonstrate compatibility with heat, salts, surfactants, or a specific emulsion system.
Test methods deserve the same scrutiny as limits. A supplier may state a viscosity range that appears aligned with an internal requirement, yet the result may have been measured at a different concentration, spindle, shear rate, or temperature. Likewise, a color range can be affected by sample preparation, illumination, or measurement scale. Ask whether the method is stated, current, and suitable for comparison. If it is not, align the methods before treating two values as equivalent.
Read the certificate of analysis as batch evidence, not a formality
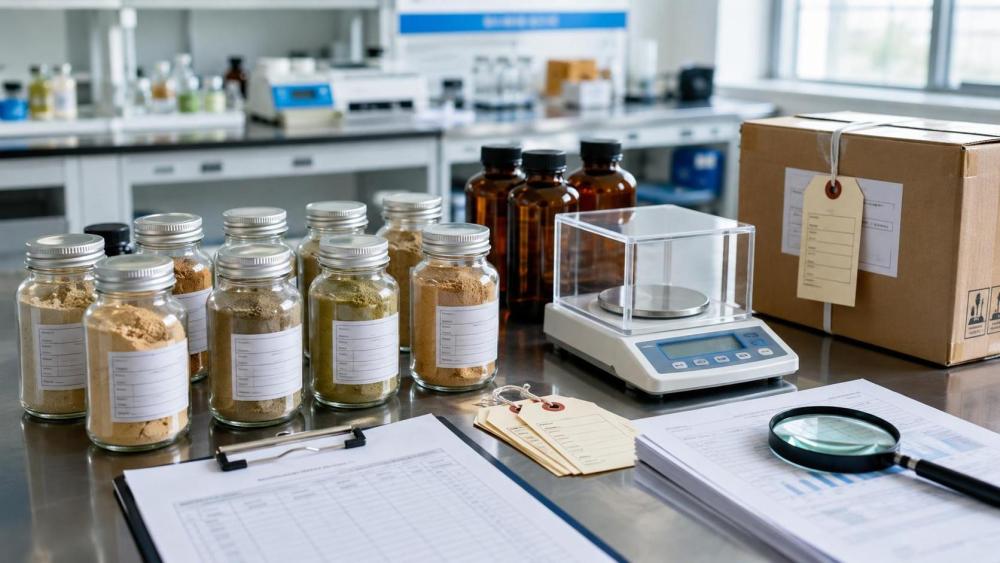
A certificate of analysis (CoA) should be traceable to a specific lot and should match the supplied material’s name, lot number, production or release date, and retest or expiry date. Compare its reported results against the current specification. A CoA that merely repeats “conforms” can be insufficient for attributes that determine product safety, identity, or clean-label status. Numeric results, test methods, and stated units provide a stronger basis for review.
Pay close attention to results reported as “not detected.” That phrase has no fixed meaning without the analytical method and reporting limit. A substance may be absent at the method’s detection threshold while still being present below that threshold. This distinction is especially relevant for pesticide residues, heavy metals, allergens, residual solvents, ethylene oxide-related concerns, preservatives, and undeclared processing contaminants. Where a claim depends on absence or a very low level, request the applicable limit of detection or quantification and confirm that the method fits the intended claim and market requirement.
Lot-to-lot consistency also matters. A single satisfactory CoA proves little about a material known to vary with harvest, geography, fermentation conditions, or extraction yield. Review a reasonable sequence of historical batch records when a material is high-risk, highly variable, or central to the finished product’s label claim. Large swings in odor, color, active content, viscosity, moisture, or microbial counts do not automatically indicate nonconformance, but they should lead to a documented explanation and an assessment of formulation impact.
Separate source claims from processing claims
“Natural,” “plant-based,” “fermented,” “non-synthetic,” and “clean label” describe different concepts. A plant source does not reveal whether the ingredient was physically milled, water-extracted, solvent-extracted, enzymatically modified, chemically reacted, bleached, deodorized, fractionated, or blended with another component. For clean-label review, the processing path often determines whether an internal policy, customer requirement, or market-specific label position remains supportable.
Request a concise process flow that identifies the input materials and the purpose of each significant processing stage. It need not expose confidential manufacturing know-how. It should clarify whether processing aids are used, whether they remain in the final material, whether solvents contact the ingredient, and whether a carrier or stabilizer is introduced after extraction or purification. A statement that no additives are “intentionally added” is narrower than a full composition declaration; it may not address residual aids, incidental carryover, or ingredients used to standardize the finished grade.
Fermented materials require particular care. A fermentation origin can be compatible with a clean-label position, but the documentation should distinguish the production organism, feedstock, downstream purification, antifoam agents, nutrient inputs of concern, and any preservative or carrier added after recovery. A declaration that an ingredient is “fermentation-derived” does not establish whether it is free from a specific allergen source, genetically modified production organism, residual protein, or processing solvent.
Resolve allergen and cross-contact statements before approval
An allergen declaration should identify intentionally added allergenic materials, relevant source materials, and the basis for any absence statement. “Allergen free” is not useful unless the document explains which allergens were assessed and whether the statement applies to formulation only or also to manufacturing-site cross-contact. Broad claims can hide important distinctions: a botanical extract might contain no added allergen while being processed on equipment that handles an allergenic carrier; a fermentation substrate may originate from an allergenic crop but not remain in detectable form after purification.
Cross-contact controls should be proportionate to the material and intended use. For dry powders, shared milling, sieving, bagging, or warehouse dust can be more relevant than for closed liquid processing. For essential oils and flavor blends, shared transfer lines and residual solvents may matter more than airborne contact. The review should identify the control point, cleaning approach, segregation practice, and whether verification relies on documented procedures, targeted testing, or both.
Do not treat a negative analytical result as a substitute for process knowledge. Testing has sampling limits, and many allergen risks arise from events that are intermittent rather than uniformly distributed. Conversely, an overly broad precautionary statement may reflect site policy rather than actual material risk. Clarification is needed before either type of statement is translated into a finished-product claim.
Confirm regulatory relevance without relying on generic letters
Regulatory declarations should name the ingredient grade, intended application category where relevant, and geographical market covered by the statement. A general letter asserting compliance with “applicable regulations” provides limited value because applicability changes with dosage, finished-product category, route of exposure, purity profile, and local labeling rules. Food, leave-on cosmetic, rinse-off cosmetic, fragrance, and household cleaning applications can require different assessments even when the ingredient’s base identity is the same.
Review whether the declaration addresses the issues that actually apply to the material: food-contact or food-use status where relevant, cosmetic restrictions, prohibited substances, restricted impurities, animal-derived inputs, genetically modified material status, irradiation status, nanomaterial status, contaminants, and country-of-origin requirements. The goal is not to collect the longest possible set of letters. It is to identify which declarations support a defined use and which are merely general corporate statements.
Where a regulatory conclusion depends on concentration in the finished formulation, preserve the calculation path. An impurity limit in the raw material may appear acceptable until the maximum use level, cumulative exposure from other ingredients, or concentration during manufacturing is considered. The reverse problem also occurs: a raw-material limitation may be stricter than needed for the intended use, causing unnecessary reformulation work because the context was not recorded.
Traceability should connect documents to physical movement
Reliable traceability links supplier lot, internal receiving record, storage condition, repackaging activity, sample identity, production batch, and final release decision. Lot numbers must remain legible and consistent across the CoA, outer packaging, delivery paperwork, retention sample label, and internal system. A mismatch that is explained verbally but not corrected in the record creates uncertainty during a complaint, recall, or customer audit.
Transportation and storage records can affect clean-label verification when materials are sensitive to heat, moisture, light, oxygen, or odor transfer. Natural pigments may fade, oils may oxidize, hydrocolloids may absorb moisture, and fragrance materials can pick up contamination from poorly controlled handling. A correct CoA at release does not prove that the material remained within specification at receipt. Review packaging type, tamper evidence, temperature requirements, humidity precautions, and the stated period after opening.
Repackaged ingredients need an especially clear chain of custody. The original supplier lot, repack date, container material, cleaning status, retained original documents, and assigned internal lot should be linked. Without that link, a valid supplier document can no longer be confidently associated with the material in use.
Use contradictions as an investigation trigger
The most useful findings often come from documents that are individually plausible but inconsistent together. A specification may identify a powder as pure extract while the allergen statement refers to a maltodextrin carrier. A technical sheet may call an ingredient water-soluble while the safety data identifies an oil-based solvent. An origin certificate may list one country while a process statement describes raw material from another region. These differences may have legitimate explanations, such as a blend made in one country from imported raw materials or a formulation change between document revisions. They should be resolved in writing before the material is released.
Maintain a controlled approval record that notes the exact documents reviewed, their revision dates, unresolved conditions, approved applications, and triggers for reassessment. Reassessment is appropriate after a site change, formulation adjustment, specification revision, new market launch, supplier notification, unexpected analytical result, complaint trend, or a change in the finished product’s clean-label wording.
Documentation verification becomes defensible when every claim can be traced to a defined material, a relevant method, a stated use, and a controlled lot history. That discipline exposes hidden carriers, weak absence claims, method mismatches, and traceability gaps before they become label, safety, or release problems.
Related News
- How to verify a cosmetic nanotechnology supplier's safety documentationOct 04, 2026Dr. Chloe Vance
- How to set a control sensory flavour standard for panel consistencySep 27, 2026Dr. Chloe Vance
- How to verify compliance documents for cosmetic active compoundsSep 19, 2026Dr. Chloe Vance
- How to verify food grade flavor formulation ingredients for safetySep 18, 2026Dr. Chloe Vance
Related News
- 00
0000-00
How to assess color stability from a natural food colors manufacturer - 00
0000-00
When does amino acid surfactant powder improve cleanser mildness? - 00
0000-00
How to verify a cosmetic nanotechnology supplier's safety documentation - 00
0000-00
How rheological thickeners for shampoo affect suspension stability - 00
0000-00
How to assess recombinant collagen ingredients for cream stability
Dr. Chloe Vance
Weekly Insights
Stay ahead with our curated technology reports delivered every Monday.